Development and Validation of an RP-HPLC Method for Estimation of Process-Related Impurities of Lacidipine in Bulk Drug and Pharmaceutical Formulations
 , Rahul Dnyaneshwar Khaire1* , Sheetal Gosavi2 , Kiran Kotade3 and Dheeraj Chechare4
1Department of Pharmaceutical Chemistry, PRES’s College of Pharmacy (For Women), Nashik, India
2Department of Pharmaceutics, PRES’s College of Pharmacy (For Women), Nashik, India
3Department of Pharmacology, PRES’s College of Pharmacy (For Women), Nashik, India
4Department of Pharmaceutics, PRES’s College of Pharmacy (D.Pharm), Nashik, IndiaCorresponding Author’s E-mail: rahuldkhaire@gmail.com
Download this article as:
ABSTRACT:Process-related impurities in active pharmaceutical ingredients (APIs) are critical quality attributes that must be identified and quantified to ensure the safety, efficacy, and regulatory compliance of pharmaceutical products. The present study aimed to synthesize a process-related impurity of Lacidipine and develop and validate a simple, sensitive, accurate, and precise reverse-phase high-performance liquid chromatography (RP-HPLC) method for its estimation in bulk drug and pharmaceutical formulations. The impurity, 3-(2-[3,5-bis(ethoxycarbonyl)-1,4-dihydropyridin-4-yl]phenyl)prop-2-enoic acid, was synthesized using the Hantzsch pyridine reaction and characterized prior to chromatographic analysis. Separation was achieved on a C18 column using an optimized methanol (60:40, v/v) mobile phase under isocratic conditions. The method was validated according to ICH Q2(R2) guidelines for specificity, linearity, precision, accuracy, sensitivity, and system suitability. The developed method exhibited excellent linearity over the concentration range of 1–6 µg/mL (R² = 0.999), with precision (%RSD <2%), satisfactory recovery (98–103%), and acceptable system suitability parameters, demonstrating reliable chromatographic performance. The validated method was successfully applied for the quantification of the process-related impurity in Lacidipine bulk drug and commercial tablet formulations, confirming its suitability for routine quality control and impurity profiling. The proposed RP-HPLC method provides a reliable, robust, and practical analytical approach for the determination of Lacidipine process-related impurities and supports regulatory requirements for pharmaceutical quality assurance.
KEYWORDS:HPLC; Impurity; Lacidipine; Method validation; ProcessValidation
Introduction
Impurities present in active pharmaceutical ingredients (APIs) have received considerable attention because they can significantly influence the safety, efficacy, and quality of pharmaceutical products.1 Regulatory authorities such as the International Council for Harmonisation (ICH) require comprehensive impurity profiling to ensure product quality and patient safety. Impurities may originate from starting materials, intermediates, by-products, degradation products, or residual solvents generated during the manufacturing process. Therefore, identification, structural characterization, and quantitative estimation of process-related impurities have become essential components of pharmaceutical quality control and regulatory submissions.2,3 Accurate impurity profiling not only ensures compliance with regulatory requirements but also minimizes the risk associated with potentially toxic impurities.4
Lacidipine is a highly lipophilic dihydropyridine calcium channel blocker widely used in the treatment of the hypertension. Several analytical methods, including RP-HPLC, HPLC-UV, and LC-MS, have been reported for the estimation of Lacidipine in bulk drugs, pharmaceutical formulations, and biological samples.5 However, most of these methods are intended for assay determination, stability studies, or pharmacokinetic investigations, with limited emphasis on the identification and quantitative estimation of process-related impurities formed during synthesis. Reliable analytical methods capable of separating Lacidipine from its process-related impurities are essential for ensuring manufacturing consistency and meeting regulatory specifications.6
To the best of our knowledge, there are very few reports describing the synthesis, characterization, and quantitative determination of the process-related impurity, 3-(2-[3,5-bis(ethoxycarbonyl)-1,4-dihydropyridin-4-yl]phenyl)prop-2-enoic acid, using a validated RP-HPLC method. The novelty of the present study lies in the successful synthesis and structural characterization of this impurity, followed by the development and validation of a simple, rapid, accurate, precise, and robust isocratic RP-HPLC method capable of achieving efficient chromatographic separation of Lacidipine and its process-related impurity in bulk drug and pharmaceutical formulations in accordance with ICH Q2(R2) guidelines. The validated method provides a practical and reliable analytical tool for routine impurity profiling, quality control, and regulatory compliance in pharmaceutical manufacturing.7
The objective of the present work was therefore to synthesize the selected process-related impurity of Lacidipine, characterize it using appropriate spectroscopic techniques, and develop and validate a robust RP-HPLC method for its quantitative determination in bulk drug and pharmaceutical formulations following ICH Q2 (R2) recommendations.8
Materials aaaand Methods
Material
The chemicals that were acquired from The Merck Chemicals Pvt. Ltd. in Nashik, Maharashtra, India, including Urea, acetone, silica gel, ethyl acetoacetate,3-(2formylphenyl)prop2enoic acid are of AR grade. We bought AR grade methanol, benzene, pyridine, ammonia, and ethyl aceto acetate from The Merck Chemicals Pvt. Ltd. in Nashik, Maharashtra, India, along with HPLC grade acetonitrile, methanol, and water.
Method0logy
Synthesis of 3-(2-[3,5bis (ethoxy carbonyl)-1, 4dihydropyridin-4-yl) phenyl) prop2enoicacid
In the presence of 10 milliliters of ethanol, 0.01 moles of 3-(2-formylphenyl) prop2enoic acid, 0.02 moles Ethyl acetoacetate, and 0.01 moles of urea were exposed to MW irradiation at 250W for two minutes at 30-second intervals. A yellow crystalline product is obtained by transferring the resulting mixture to ice-cold water. Title chemical was obtained by filtering and re crystallizing alcohol.9
HPLC Method Development and Validation
The method developed with LC20AD Shimadzu, Japan, which has a UV-Vis detector and a C18 column measuring 25×0.6 cm and 1.0 ml/min rate of flow at 234nm wavelength. 1, 4-DHP was measured from both bulk drug and the formulation using the HPLC method. The mobile phase used to create the chromatogram is a 60:40 ratio of methanol to water. The process involved validating the synthesized molecule and examining a number of parameters in accordance with ICH criteria Q2(R2).10,11
Analytical Method Validation
For identification, an appropriate analytical technique was created and verified. At different phases of development, new medication development necessitates the production of significant and trustworthy analytical data.12
Preparation of Mobile phase
The solvents’ polarity and non-polarity were taken into consideration when choosing the mobile phase. After choosing a mobile phase as 60:40 methanol: water, the mixture was agitated 10 to 15 minute and filtered through a membrane filter (0.45 μ) to eliminate degassing.13
Preparation of Stock solution standard
The standard procedure, which involves carefully weighing 10mg of the molecule synthesized on an analytical scale and mobile phase to reach a volume up to 100ml, was followed to create the stock solution. For the HPLC technique validation, the sample was produced at concentrations ranging from 1 to 6 ppm.14,15
Preparation of sample solution (Formulation)
Two separate batches of the commercial formulation of Lacidipine, each containing 100 ppm, were made in volumetric flask of 100 ml along with the stock solution of bulk Lacidipine. It is necessary to dissolve 10mg of the test sample in diluents 100 ml. One milliliter of this solution of stock was diluted to ten milliliters to make a 10 ppm stock solution. For the tablet formulation, twenty tablets were crushed from each of the two tablet batches. The powder drug equal to 10 mg used to that of standard. They were created by first diluting 10 ml of the standard test solution to 0.1 ml, 0.2 ml, and so on, and then diluting it even further to 1 ppm, 2 ppm, and so on.16
Quantitation of Impurity
The 0.1% ICH limit for impurities in novel medicinal compounds was compared to the total impurity concentration of the synthesized compound in Lacidipine bulk and formulations.17
Results
HPLC Chromatograph of Lacidipine
Figure 1 shows the graph and data for HPLC chromatogram and TheRT of Lacidipine found to be 9.5min.
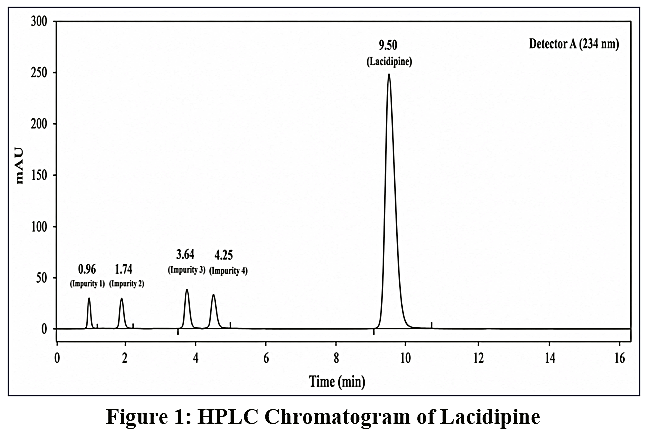 |
Figure 1: HPLC Chromatogram of Lacidipine Click here to View Figure |
Synthesized compounds HPLC Chromatogram-
10.5Min was the retention time of impurity and as depicted in Figure 2 showing single peak in the graph.
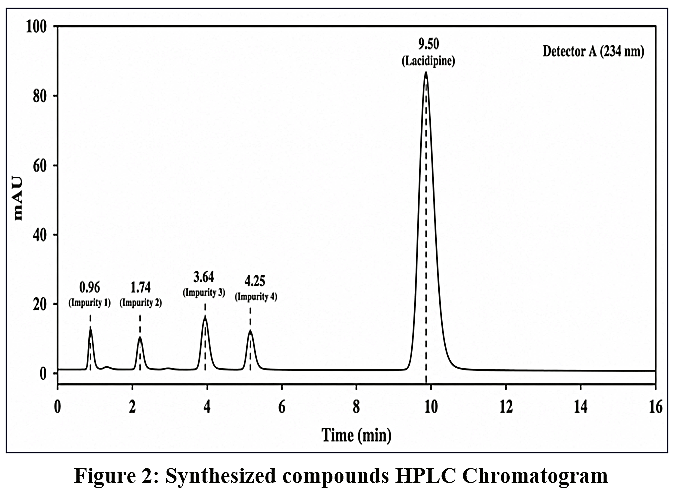 |
Figure 2: Synthesized compounds HPLC Chromatogram Click here to View Figure |
HPLC chromatogram of the combination
Lacidipine and synthesized molecule were shown to have retention times of 9.60 and 10.49 minutes, respectively, in the lab mixture as shown in Figure 3.
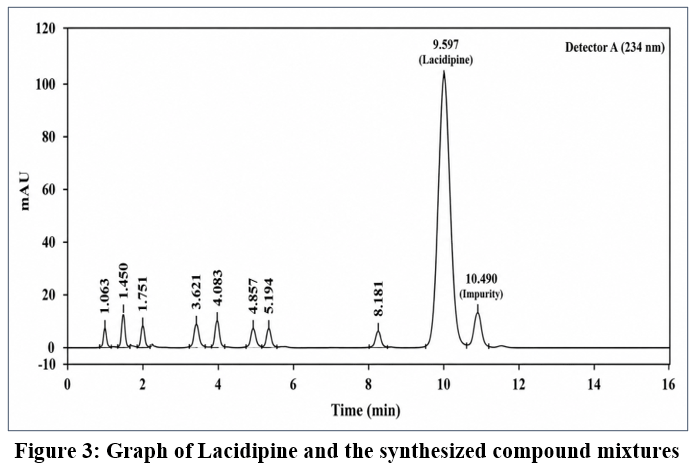 |
Figure 3: Graph of Lacidipine and the synthesized compound mixtures Click here to View Figure |
Tablets HPLC Chromatogram –
As shown in Figure 4 the 9.6 minutes was the Lacidipine tablet’s retention time.
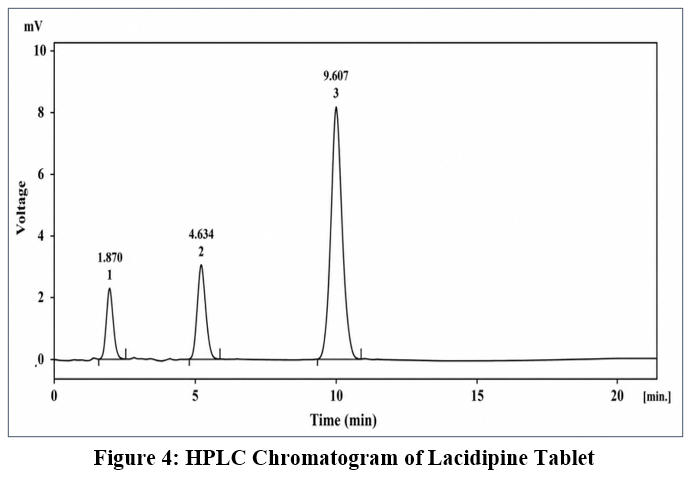 |
Figure 4: HPLC Chromatogram of Lacidipine Tablet Click here to View Figure |
Tablet and synthetic chemical mixture HPLC chromatogram
As shown in Figure 5, the retention times for the synthesized chemical and Lacidipine tablet in the lab mixture were 9.60 and 10.58 minutes, respectively.
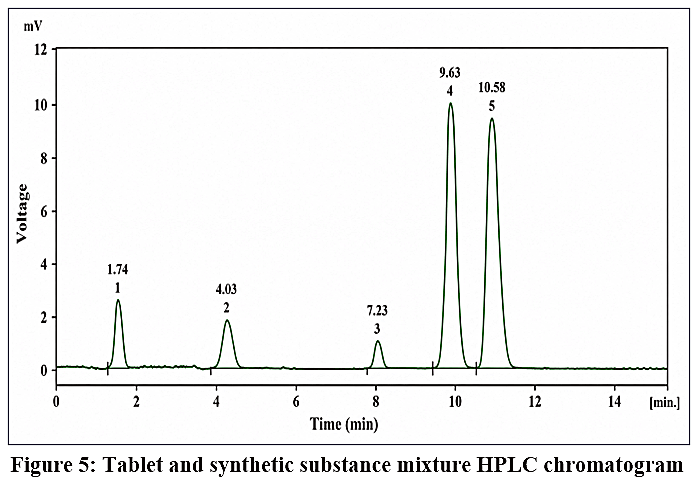 |
Figure 5: Tablet and synthetic substance mixture HPLC chromatogram Click here to View Figure |
Optimized chromatographic condition-
A Shimadzu HPLC system with an ARP-C18 column was used for the analysis. The mobile phase was supplied at a flow rate of one milliliter per minute and included methanol and water in a 60:40 (v/v) ratio. The injection volume was 20 µL, and the detection wavelength was set at 234 nm. With a retention period of roughly 10.5 minutes, the separation was performed at room temperature.18
Linearity
The linearity was assessed at six distinct concentration levels, ranging from 1 to 6 µg/ml for intermediate, as shown in Table 1 Linearity result and Figure 6 shows that the 0.999 correlation coefficient (R2) and it was determined to be linear.
Table 1: Linearity result
|
Sr. No |
Concentration |
Area |
|
1 |
1 ppm |
124 |
|
2 |
2 ppm |
215 |
|
3 |
3 ppm |
312 |
|
4 |
4 ppm |
419 |
|
5 |
5 ppm |
513 |
|
6 |
6 ppm |
618 |
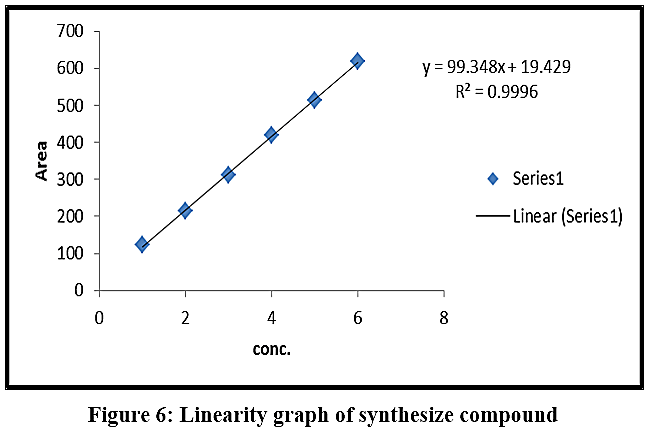 |
Figure 6: Linearity graph of synthesize compound Click here to View Figure |
Precision
As shown in the Table 2 Precision by HPLC. The intermediate’s accuracy was consistent with its chromatogram area and was assessed for a range of repeated concentrations of 4µg/ml. According to the findings, the SD was 1.331 and RSD was 0.317.
Table 2: Precision study by HPLC
|
Sr No |
Area (4 ppm conc) |
Mean |
Std. Deviation |
% Relative Std Dev. |
|
1. |
419.19 |
419.89 |
1.331 |
0.317 |
|
2. |
418.98 |
|||
|
3. |
421.90 |
|||
|
4. |
419.90 |
|||
|
5. |
418.39 |
|||
|
6. |
421.01 |
Summary of Precision Studies
The precision was evaluated through repeatability, intraday, Interday, robustness, and ruggedness studies. The SD and %RSD determined for each parameter. The SD and %RSD for precision were found to be 1.332 and 0.317. For Intraday, the SD was 2.250 and %RSD was 0.539, while for Interday precision, the SD and %RSD were 1.760 and 0.414, respectively. The robustness study showed SD and %RSD values of 3.122 and 0.395, respectively, and the ruggedness study showed values of 1.436 and 0.341, respectively. All %RSD values were found to be ≤ 2.
Accuracy
The accuracy was checked by observing recovery studies at three conc. levels of 50%, 75%, and 100%. The results demonstrated excellent recovery for both the bulk drug and tablet formulations.19
For the bulk drug, the percentage recoveries at 50%, 75%, and 100% levels were 98.18%, 99.07%, and 99.89%, respectively, with a mean recovery of 99.04%, standard deviation (SD) of 0.852, and %RSD of 0.863.
For Tablet I, recoveries at 50%, 75%, and 100% were 99.22%, 101.30%, and 103.79%, respectively, giving a mean recovery of 101.43%, SD of 2.288, and %RSD of 2.255.
For Tablet II, recoveries were 99.25%, 101.68%, and 103.13% at 50%, 75%, and 100% levels, respectively, with a mean recovery of 101.30%, SD of 1.969, and %RSD of 1.934.
All %RSD values were within acceptable limits of ≤ 2%
Limit of detection- By HPLC, the LOD was 135.2 ng.
Limit of quantitation- By HPLC, the LOD was 446.2 ng.
System Suitability Parameters
The retention time of the analyte was 10.59 min, with a relative standard deviation (%RSD) within 1%, confirming excellent reproducibility. The theoretical plate count (N) was observed to be 10,224, which is well above the required limit of N ≥ 2000, indicating satisfactory column efficiency. The resolution (R) between the peaks was 6.36, exceeding the minimum acceptance criterion of R ≥ 2, demonstrating good separation between analyte peaks.20
Quantitation of Synthesized Compound
Lacidipine’s process-related impurities in tablets and bulk were quantified. Tablets I and II had impurity levels of 0.28% and 0.33%, respectively, and no impurity was observed in large quantities. When the impurity content in tablet formulations exceeds 0.1%, as defined by the ICH limit, it is considered.
Discussion
The present study successfully developed and validated a simple, accurate, and robust RP-HPLC method for the quantification of a process-related impurity of Lacidipine in bulk drug and pharmaceutical formulations. The developed method achieved satisfactory chromatographic separation with good peak resolution and demonstrated excellent linearity (R² = 0.999), precision (%RSD < 2%), and accuracy (98–103% recovery), confirming its reliability for routine analysis. Compared with previously reported RP-HPLC methods, which primarily focus on the assay of Lacidipine or stability studies, the present method specifically addresses the determination of a synthesized process-related impurity, thereby providing an additional tool for impurity profiling and regulatory quality assessment. The impurity levels detected in commercial formulations highlight the importance of routine monitoring to ensure compliance with ICH requirements. Although the method was validated successfully, further studies involving additional process- and degradation-related impurities and LC–MS-based confirmation may broaden its applicability. Overall, the developed method is suitable for routine quality control and pharmaceutical impurity profiling of Lacidipine.
Conclusion
The present study successfully synthesized and characterized the process-related impurity, 3-(2-[3,5-bis(ethoxycarbonyl)-1,4-dihydropyridin-4-yl]phenyl)prop-2-enoic acid, of Lacidipine and developed a simple, rapid, and reliable isocratic RP-HPLC method for its quantitative determination in bulk drug and pharmaceutical formulations. The developed method achieved efficient chromatographic separation of Lacidipine and its process-related impurity with excellent specificity and was validated in accordance with ICH Q2(R2) guidelines. Validation results demonstrated excellent linearity (R² = 0.999), satisfactory accuracy (98–103% recovery), high precision (%RSD < 2%), and acceptable system suitability parameters, confirming the robustness and reproducibility of the method. The method was successfully applied for impurity profiling in commercial formulations, demonstrating its suitability for routine quality control and regulatory compliance. The proposed RP-HPLC method provides a simple, sensitive, accurate, and cost-effective analytical tool for the determination of Lacidipine process-related impurities and can be effectively employed in pharmaceutical industries for routine analysis, quality assurance, and impurity monitoring.
Acknowledgement
The authors gratefully acknowledge the Department of Pharmaceutical Chemistry, PRES’s College of Pharmacy (For Women), Nashik, India, for providing the necessary laboratory facilities, analytical infrastructure, and technical support required to carry out this research work.
Funding Sources
The author(s) received no financial support for the research, authorship, and/or publication of this article.
Conflict of Interest
The authors do not have any conflict of interest.
Data Availability Statement
This statement does not apply to this article.
Ethics Statement
This research did not involve human participants, animal subjects, or any material that requires ethical approval.
Informed Consent Statement
This study did not involve human participants, and therefore, informed consent was not required.
Clinical Trial Registration
This research does not involve any clinical trials.
Permission to Reproduce Material from other Sources
Not Applicable.
Author Contributions
- Kiran Dhamak: Conceptualization, methodology, investigation, data curation, writing—original draft preparation.
- Rahul Dnyaneshwar Khaire: Supervision, Validation, formal analysis, writing—review and editing, project administration.
- Sheetal Gosavi: Methodology, experimental work, data analysis.
- Kiran Kotade: Resources, software, data interpretation.
- Dheeraj Chechare: Visualization, validation, review of manuscript.
References:
- Ramesh G, Vamshi Vishnu Y, Chinna Reddy P, et al. Development of high performance liquid chromatography method for lacidipine in rabbit serum: application to pharmacokinetic study. Anal Chim Acta. 2009;632:278-283.
CrossRef - Bendre SD, Jaiswal S, Jain SK, Vengurlekar S. Analytical method development of process related impurities of drug lacidipine by using RP-HPLC techniques. J Neonatal Surg.
- Khaire RD, Bhangale CJ, Pagare RV, Chothave SJ, Gosavi SB, Jagtap BT. Stability indicating RP-HPLC method development and validation for gatifloxacin and prednisolone acetate. Rasayan J Chem. 2025;18(4).
CrossRef - Vinod M, Vinayak M, Rahul K, Pankaj P. Development and validation of a HPLC method for the determination of lacidipine in pure form and in pharmaceutical dosage form. Malays J Anal Sci. 2012;16:213-219.
- Khaire RD, Pawar PY. A review on development of stability indicating analytical methods for drugs in bulk and pharmaceutical dosage form. Asian J Pharm Anal. 2022;12(4):261-263.
CrossRef - Shantha Kumar PT, Siva Kumar U, Nagaraju PT. Int J Pharm Sci Rev Res. 2010;5:14-18.
- Panda SS, Bera RKVV, Acharjya SK, Sahoo P, Beg S. Analytical lifecycle management approach: application to development of a reliable LC method for estimation of lacidipine. Sep Sci Plus. 2019;2:18-25.
CrossRef - Khaire RD, Gaware VM, Dhamak KB, Jagtap BT, Gaikar MT, Kunde VD. Validated stability-indicating HPTLC method for quantification of nortriptyline. Rasayan J Chem. 2026;19(2):801-806.
CrossRef - Carling RS, Degg TJ, Allen KR, Bax NDS, Barth JH. Evaluation of whole blood serotonin and plasma and urine 5-hydroxyindole acetic acid in diagnosis of carcinoid disease. Ann Clin Biochem. 2002;39:577-582.
CrossRef - Maasma Shaik, Sunandamma D, Reddy CS. Development and validation of RP-HPLC assay method for determination of lacidipine in tablets. J Pharm Sci Res. 2019;11:2204-2207.
- Moorthi C, M A, Prezzy E, Sumithira G. Spectrophotometric determination of lacidipine in bulk and tablet dosage form. Journal of Applied Pharmaceutical Science. 2011; 1(5):209-13.
- Dongala T, Katari NK, Palakurthi AK, Jonnalagadda SB. Development and validation of a generic RP-HPLC PDA method for the simultaneous separation and quantification of active ingredients in cold and cough medicines. Biomed Chromatogr. 2019;33:e4641.
CrossRef - Zagalo DM, Sousa J, Simões S. Quality by design (QbD) approach in marketing authorization procedures of non-biological complex drugs: a critical evaluation. Eur J Pharm Biopharm. 2022;178:1-24.
CrossRef - Agrawal D, Dahiya M, Wakode S, Singh GP, Shiv J. Method development and validation of stability indicating HPLC assay for the determination of tenofovir alafenamide fumarate in tablet formulation. International Journal of Pharmaceutical Sciences and Nanotechnology. 2020;13(6):5226-33.
CrossRef - Jagdale AS, Pendbhaje NS, Nirmal RV, Bachhav PM, Sumbre DB. Development and validation of RP-HPLC method for estimation of brexpiprazole in its bulk and tablet dosage form using quality by design approach. Future J Pharm Sci. 2021;7:142.
CrossRef - Khalid MB, Shahzad F, Siddiqui MR, et al. Comparative efficacy and safety of irreversible (rasagiline) and reversible (safinamide) monoamine oxidase inhibitors as add-on therapy for Parkinson’s disease. J Neurol. 2025;272:486.
CrossRef - Bhosale M, Khaire R, Wagh M, Chechare D, Thombare Y. Analytical Method Development and Validation for Safinamide Mesylate Using RP-HPLC with Quality by Design Approach. Biosci Biotech Res Asia 2026;23(2).
CrossRef - Mohapatra D, Kumar DN, Shreya S, et al. Quality by design–based development and optimization of fourth-generation ternary solid dispersion of standardized Piper longum extract for melanoma therapy. Drug Deliv Transl Res. 2023;13:3094-3131.
CrossRef - Khaire RD, Gosavi SB, Chothave SJ, Pagare RV, Chechare DD. Stability indicating method development and validation of Dolutegravir by RP-HPLC. Intern J Zool Invest. 2026;12(1):1076-1083.
CrossRef - Desai R, Dhumal P. RP-HPLC Methodology for Olanzapine Impurity Analysis: Advances in Development and Validation. International Journal of Phytochemistry and ChemAnalysis. 2024; 1(01):14-25.
Abbreviations
RP-HPLC: Reversed-Phase High-Performance Liquid Chromatography
RSD: Relative Standard Deviation
RT: Retention Time
SD: Standard Deviation
TP: Theoretical Plates
UV: Ultraviolet
% RSD: Percentage Relative Standard Deviation
Accepted on: 13-08-2026
Second Review by: Dr. Ramya Sri
Final Approval by: Dr. Muhammad Hamayun